Oral Presentation Crystal36-AXAA Conference 2026
Optimizing the connectivity of protein conformations to untangle ensemble refinement (143478)
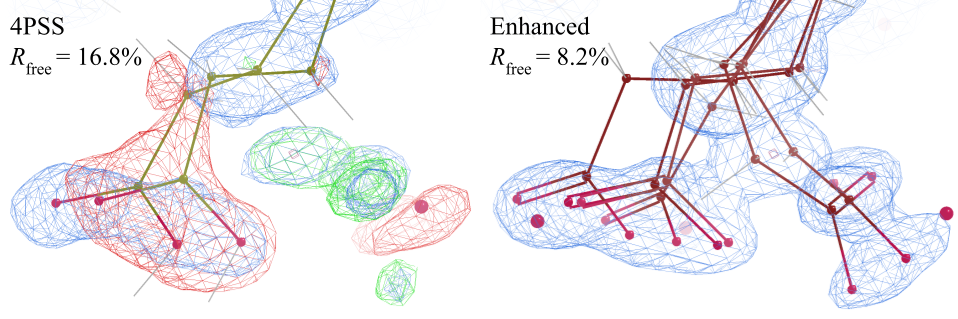
Protein crystallography captures the diffraction signal of the conformational ensemble of proteins contained within a crystal. As such, modelling proteins in multiple conformations may offer particularly realistic representations of collected data. In practice, multiconformer refinement produces only minor improvements in agreement with experimental data (Rfree). It was recently shown[1] that multiconformer models are universally trapped - rather than fitting the electron density with the collection of model conformations harmoniously, refinement algorithms strain each model conformation to fit the electron density in its immediate vicinity. Here, we demonstrate that this trap may be escaped by formulating, as an integer linear programming problem, the construction of low-energy conformations from the coordinates of atom alternate locations (altlocs). This approach forms the core of an automated refinement method that we present as a solution to the "Untangle Challenge"[1] - created to solicit solutions to this specific problem. Combined with other steps to minimize errors specific to multiconformer models, we present a 6-conformation model for a deposited 0.85 Å dataset of DHFR[2] (PDB ids: 4PTH, 4PSS) with substantially reduced Rwork/Rfree values of 6.3%/8.2%.
- Hopkins MS, Terwilliger TC, Afonine PV, Ginn HM, Holton JM. The Untangle Challenge for accurate ensemble models. bioRxiv 2026.02.21.706873, (2026), doi: 10.64898/2026.02.21.706873
- Keedy D et al. Crystal Cryocooling Distorts Conformational Heterogeneity in a Model Michaelis Complex of DHFR. Structure, (2014), 22, 899-910, doi: 10.1016/j.str.2014.04.016